EU MDR Article 117: What Pharma Sponsors Need From Injection Device Suppliers
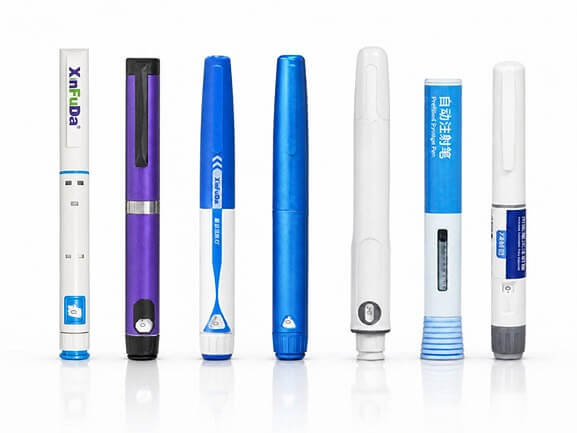
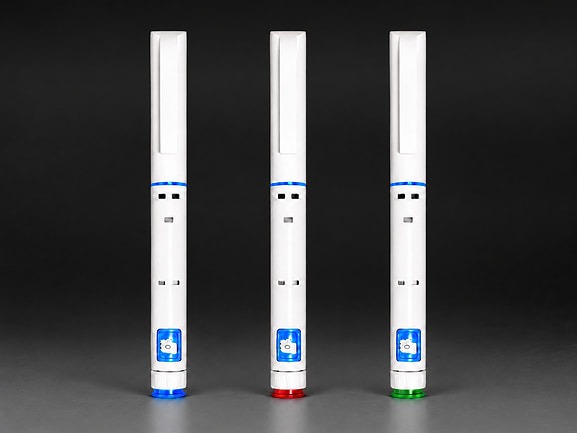
injecion pen types
When a medicinal product and delivery device form one integral product, the European marketing authorisation application may need evidence addressing the device part under Article 117 of Regulation (EU) 2017/745.
The European Medicines Agency lists prefilled syringes and prefilled pens as examples of integral drug-device combinations. EMA explains that the application should include a CE certificate for the device part or, where applicable, a notified body opinion on conformity with the relevant General Safety and Performance Requirements.
For injection pen and autoinjector programs, Article 117 is not a document to request from the supplier at the end. Device evidence, responsibilities and change control should be defined early.
First Confirm the Configuration
| Configuration | Example | Starting Point |
| Integral product | Prefilled pen or PFS autoinjector | Medicinal product pathway; Article 117 may apply to the device part |
| | Co-packaged | Medicine and separate device in one pack | Device generally needs applicable conformity evidence |
| | Separately obtained | Reusable pen used with replaceable cartridges | The pen may be regulated as a separate medical device |
Documentation to Discuss With the Supplier
1.Device Configuration
The supplier should clearly identify the platform, components, drawings, materials and primary container interfaces included in the assessed version.
2.Risk Management
Relevant risks may include incorrect dose setting, incomplete delivery, unintended activation, container damage and use error. Risk controls should connect to verification evidence.
3. Verification
Depending on the product, evidence may cover dose delivery, activation, operating force, injection time, container fit, environmental conditioning and packaging.
4.Materials and Manufacturing
The sponsor needs enough information to understand critical materials, manufacturing controls, traceability and changes that could affect the device.
5.Labeling and Instructions
The final label, carton and instructions should match the assessed configuration and intended use.
6.Change Control
The agreement should define which material, component, tooling, site or process changes require notification and impact assessment.
Article 117 Readiness Checklist
- Is the product integral, co-packaged or separately obtained?
- Which device classification would apply if the device were marketed separately?
- Who prepares the GSPR mapping?
- Is the assessed device identical to the intended commercial configuration?
- Are cartridge or PFS interfaces controlled?
- Which performance and usability tests support the final configuration?
- How will supplier changes be communicated?
- When will the notified body be engaged?
Why the Primary Container Matters
In a prefilled pen or autoinjector, the primary container is both packaging and a functional part of the delivery system.
Cartridge or PFS dimensions can affect assembly, activation and dose delivery. Formulation viscosity can influence drive force and injection time. A different stopper, plunger or needle system may require additional assessment.
Evidence generated with one container should not automatically be assumed to support another materially different container.
Platform Evidence Versus Product-Specific Evidence
An established platform can provide useful design and manufacturing history. However, the sponsor still needs product-specific review of the actual container, dose, formulation, users, labeling and packaging.
This distinction prevents two mistakes: rebuilding every element from the beginning or assuming platform history alone supports the final medicinal product.
What Pharma Companies Should Ask Before Selection
1. Which platform evidence can be provided for assessment?
2. How will container compatibility be evaluated?
3. Which features can be customized without changing the architecture?
4. How are device versions and manufacturing changes controlled?
5. What support is available for samples, technical questions and notified body responses?
Frequently Asked Questions
1.Does Article 117 apply to every reusable injection pen?
Not automatically. The product configuration and intended use determine whether the device is integral, co-packaged or supplied separately.
2.Is platform documentation enough?
Not by itself. The final drug, container, dose, labeling and use conditions require product-specific assessment.
3.When should notified body planning begin?
Before the evidence package and submission timeline are fixed, so documentation gaps can be resolved without late redesign.


Copyright © Shijiazhuang Xinfuda Medical Packaging Co., Ltd. All Rights
MAKE AN ENQUIRY